Perfusion device safety extends beyond the pre-bypass checklist. Teams also need a practical way to recognize equipment alerts, identify affected supplies, and communicate required actions before a problem reaches the bedside. The International Perfusion Association’s Perfusion & ECMO Device Safety resource brings FDA recall records and reported device problems into a searchable starting point for that work.
The resource is intended to support informed review. Its value depends on what happens after a search: opening the original notice, matching the affected product to local inventory, and confirming that the appropriate institutional response is completed.
A Focused Resource for Perfusion and ECMO
The page provides separate views for FDA recall records and Manufacturer and User Facility Device Experience (MAUDE) reports. Users can narrow searches by equipment category, product or catalog number, manufacturer, and date. Coverage includes cardiopulmonary bypass and ECMO equipment, oxygenators, circuits, cannulas, heater-coolers, monitoring, blood-management systems, coagulation testing, and mechanical circulatory support.
Selected official notices are also highlighted. These summaries provide a route to the original FDA material; the current official notice remains the source for affected models, lots, and required actions. The page displays review and data-update information so readers can judge the age of the material they are viewing.
Recall Records and MAUDE Reports Answer Different Questions
A recall notice identifies a product issue and the associated corrective action. FDA also publishes early alerts while it is gathering information about potentially serious device problems. A notice may describe removal, correction, or updated use instructions; the required response must be read in the specific notice rather than inferred from the headline.
MAUDE contains reports of suspected device-associated adverse events and malfunctions. Submission does not establish that the device caused an event. Reports may be incomplete, inaccurate, delayed, or unverified, and the database lacks a reliable denominator for device use. Report totals therefore cannot establish failure rates or support comparisons of safety between products. Multiple reports and supplemental information also complicate interpretation.
For a perfusion service, the practical distinction is straightforward: a recall requires review of the stated action and affected inventory, while an individual adverse-event report is a signal that needs context and investigation.
Turning a Notice Into a Local Safety Check
The following is a suggested departmental workflow, rather than a protocol tested by the IPA resource:
- Confirm the source. Open the FDA or manufacturer notice and check for revisions, related notices, and the date of the manufacturer’s communication.
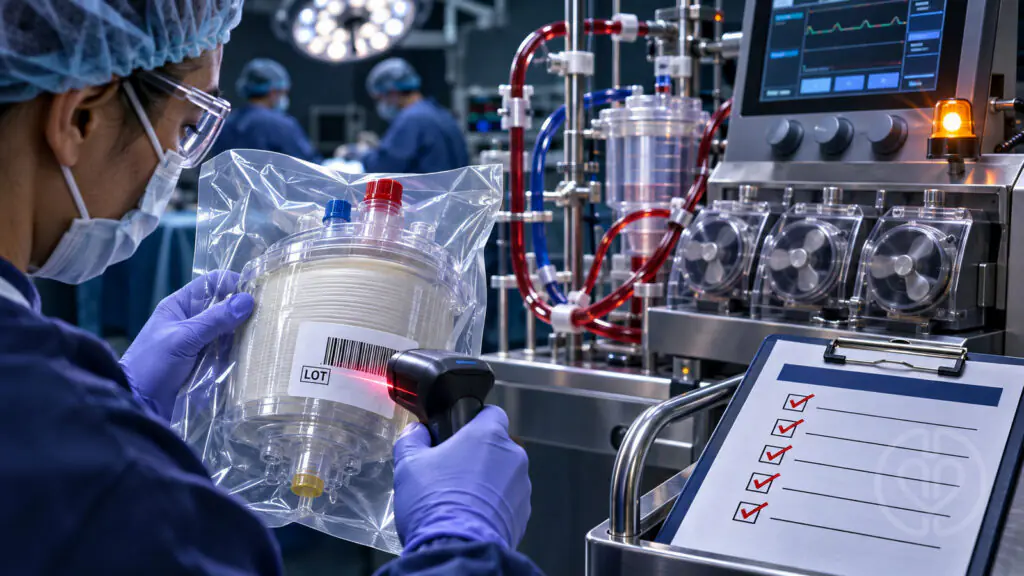
- Identify the exact equipment. Compare the catalog number, model, unique device identifier, lot or serial number, and software version where relevant. A familiar product name alone is insufficient to establish that local stock is affected.
- Locate supplies across the service. Consider the operating room, ECMO storage, emergency carts, transport equipment, loaned consoles, and reserve inventory. Assign responsibility for checking each location.
- Carry out the stated response. Work with supply chain, biomedical engineering, clinical leadership, and the manufacturer as appropriate. Document any required quarantine, return, service, or instruction update under institutional procedures.
- Close the communication loop. Record who reviewed the notice, whether affected products were found, what action was completed, and what remains outstanding. Include relevant changes in handovers and staff briefings.
Prepare for the Equipment You May Need Next
Safety review is also an opportunity to examine backup readiness. If an affected component is removed from inventory, the team should know which approved alternative is available, where it is stored, and whether staff are familiar with it. An alternative that requires different connectors, accessories, or setup steps deserves attention before an emergency.
A short recurring review can make this work manageable. One designated reviewer can screen notices, while the person responsible for each equipment group confirms local applicability. Keeping a shared action log helps distinguish an alert that has been read from one whose required response has been completed.
Limits and Responsible Use
The IPA resource uses selected FDA product codes and search terms; it is not guaranteed to retrieve every relevant notice or report. Data publication and review occur on different schedules. An empty search result should not be treated as confirmation that equipment has no known safety concerns.
This article describes an educational resource and proposes a practical review workflow. It does not demonstrate that use of the page reduces complications or replace device instructions, manufacturer communications, institutional reporting pathways, or clinical judgment.
Explore the Device Safety Resource
Open Perfusion & ECMO Device Safety to search records and follow links to the original notices. Check the page’s displayed update information each time you use it.
Sources
- International Perfusion Association. Perfusion & ECMO Device Safety. Accessed November 6, 2026.
- U.S. Food and Drug Administration. Medical Device Recalls and Early Alerts. Accessed November 6, 2026.
- U.S. Food and Drug Administration. About the MAUDE Database. Accessed November 6, 2026.
Disclosure: This overview describes IPA’s own educational resource. The cover is an AI-generated editorial illustration and does not depict a specific recalled device or a validated circuit configuration.